
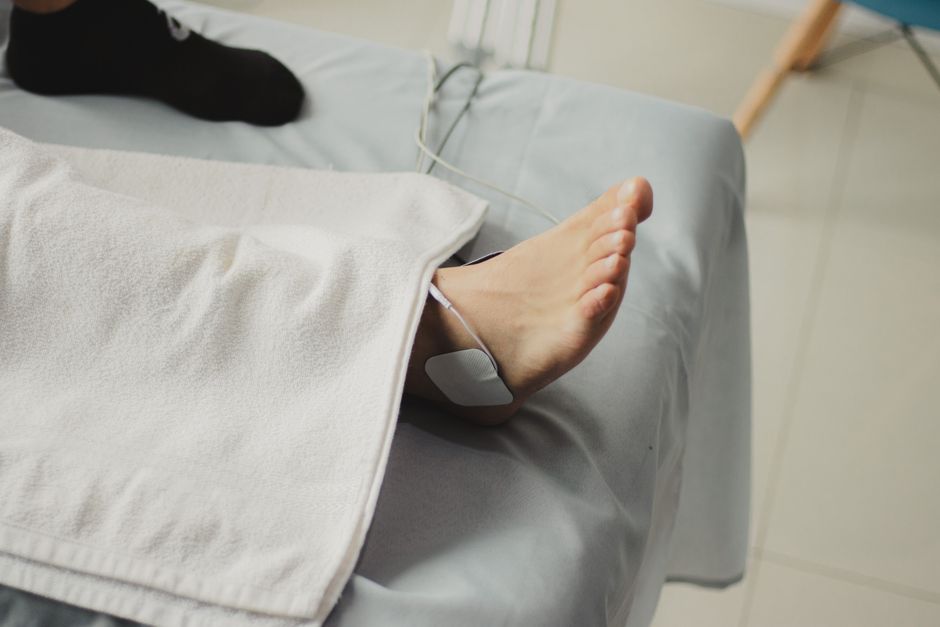
Charcot-Marie-Tooth disease is one of the most common inherited nerve disorders, affecting roughly 1 in 2,500 people. It damages the peripheral nerves — the ones connecting your brain and spinal cord to your muscles and skin — and slowly weakens the muscles in the feet, lower legs, and hands. It is not fatal, it does not shorten life expectancy, and it is not the same as muscular dystrophy or multiple sclerosis, though people often confuse the three.
What Is Charcot Marie Tooth Disease?
CMT, as it is usually called, is a group of inherited conditions that affect the peripheral nerves. These nerves carry signals from your brain and spinal cord out to your muscles, and carry sensation back the other way.
In CMT, the nerves themselves gradually break down. The myelin sheath — the protective coating around many nerves — may be damaged, or the nerve fiber itself (the axon) may degenerate. Different genetic types affect different parts of the nerve. The result is the same: weaker signals getting through.
It is named after the three physicians who described it independently in 1886: Jean-Martin Charcot and Pierre Marie in France, and Howard Henry Tooth in England. The name is historical, not descriptive.
One thing worth clarifying: CMT is a nerve disease, not a muscle disease. The muscles weaken because the nerves supplying them are failing. That distinction matters for how the condition is managed and for understanding why certain symptoms appear.
What Causes Charcot-Marie-Tooth Disease?
CMT is genetic. It is caused by mutations in genes that control how peripheral nerves are built and maintained. More than 100 genes have been linked to different forms of the disease.
Most types are inherited in one of three patterns:
- Autosomal dominant: One copy of the altered gene from one parent is enough to cause the disease. This is the most common pattern.
- Autosomal recessive: Both parents must pass on an altered copy. Each parent is usually an unaffected carrier.
- X-linked: The altered gene sits on the X chromosome. This pattern tends to affect males more severely than females.
CMT1A, the most common subtype, is caused by a duplication of the PMP22 gene and accounts for a large share of all cases. It follows an autosomal dominant pattern.
A significant number of cases — estimates vary by study — arise from new mutations with no family history at all. So a person with CMT does not necessarily have an affected parent.
What Are the Symptoms of Charcot-Marie-Tooth Disease?
CMT usually starts gradually, often in childhood or adolescence, though some forms begin in adulthood. Symptoms build slowly over years, and the pattern is fairly consistent across most types.
The earliest and most common signs involve the feet and lower legs:
- High arches (pes cavus) or, less often, flat feet
- Weakness in the muscles that lift the foot, leading to foot drop and tripping
- Hammertoes and curled toes
- Reduced sensation in the feet and lower legs
- Cold feet or numbness
- Muscle wasting in the lower legs, giving the calves a thin, “inverted champagne bottle” appearance
Later, the hands are often affected. Fine motor tasks — buttoning a shirt, opening jars, writing — become harder. Some people develop tremor in the hands.
Balance problems are common because the brain relies on sensory feedback from the feet to stay steady. When that feedback fades, people compensate by watching their feet or widening their stance.
Pain is variable. Some people have cramping or nerve pain; others have little discomfort despite noticeable weakness. Fatigue is frequently reported.
Symptoms usually progress slowly. Most people with CMT remain able to walk, often with aids such as braces or a cane later in life. A minority need a wheelchair. The rate of progression varies widely, even within the same family.
How Is Charcot-Marie-Tooth Disease Diagnosed?
Diagnosis starts with a neurological exam. A doctor checks muscle strength, reflexes, sensation, and the shape of the feet. Reduced or absent ankle reflexes are a classic finding.
Nerve conduction studies and electromyography (EMG) measure how well electrical signals travel along the nerves. In CMT, nerve conduction velocity is often slowed, and the pattern helps distinguish broad categories of the disease.
Genetic testing can confirm the specific type. It is increasingly used early because identifying the exact genetic cause matters for family planning and, in some cases, for clinical trials.
Family history is important, but as noted, its absence does not rule out CMT. A genetic counselor can help interpret results and discuss inheritance risks.
How Is Charcot-Marie-Tooth Disease Treated?
There is no cure for CMT, and no medication has been approved to slow or reverse it. Treatment focuses on managing symptoms, preserving function, and preventing complications.
Physical and occupational therapy are central. Stretching and strengthening exercises help maintain range of motion and muscle function. Ankle-foot orthoses (braces) are commonly used to correct foot drop and improve walking safety.
Orthopedic evaluation may be recommended for foot deformities. Surgery is sometimes considered for severe structural problems, though outcomes vary and it is not appropriate for everyone.
For nerve pain, some clinicians prescribe medications used for other types of neuropathic pain, such as certain antidepressants or anticonvulsants. These are used off-label for CMT-related pain, and evidence specific to CMT is limited.
Regular foot care matters. Because sensation is reduced, small injuries, blisters, or pressure sores can go unnoticed and worsen. People with CMT should inspect their feet regularly and wear well-fitting shoes.
Genetic counseling is recommended for anyone considering children, given the inherited nature of the disease.
What Is the Outlook for Someone With CMT?
Most people with CMT live a normal lifespan. The disease is progressive but usually slow, and many people adapt well with therapy, braces, and lifestyle adjustments.
Function varies enormously between types and between individuals. Some people have mild symptoms that barely affect daily life. Others experience significant disability over decades. Predicting the course for a specific person is difficult, even with genetic testing.
Regular follow-up with a neurologist or a multidisciplinary clinic can help catch problems early — such as falls, foot injuries, or hand weakness — before they become serious.
Frequently Asked Questions
Is Charcot-Marie-Tooth disease fatal?
No. CMT does not shorten life expectancy in most cases and is not considered a fatal disease. Rare severe forms exist, but the vast majority of people with CMT live a normal lifespan.
Can Charcot-Marie-Tooth disease be cured?
There is currently no cure and no approved treatment that slows or reverses the nerve damage. Management focuses on therapy, braces, foot care, and symptom control.
Is CMT the same as muscular dystrophy?
No. CMT is a disease of the peripheral nerves, while muscular dystrophy primarily affects the muscle tissue itself. They are distinct conditions with different genetic causes and management.
Can you prevent Charcot-Marie-Tooth disease?
You cannot prevent an inherited genetic condition, but genetic counseling can help families understand their risk. Prenatal and preimplantation genetic testing are options some families consider.
